Economic Operators Under the MDR: Manufacturer, Importer, Distributor and Authorised Representative Obligations
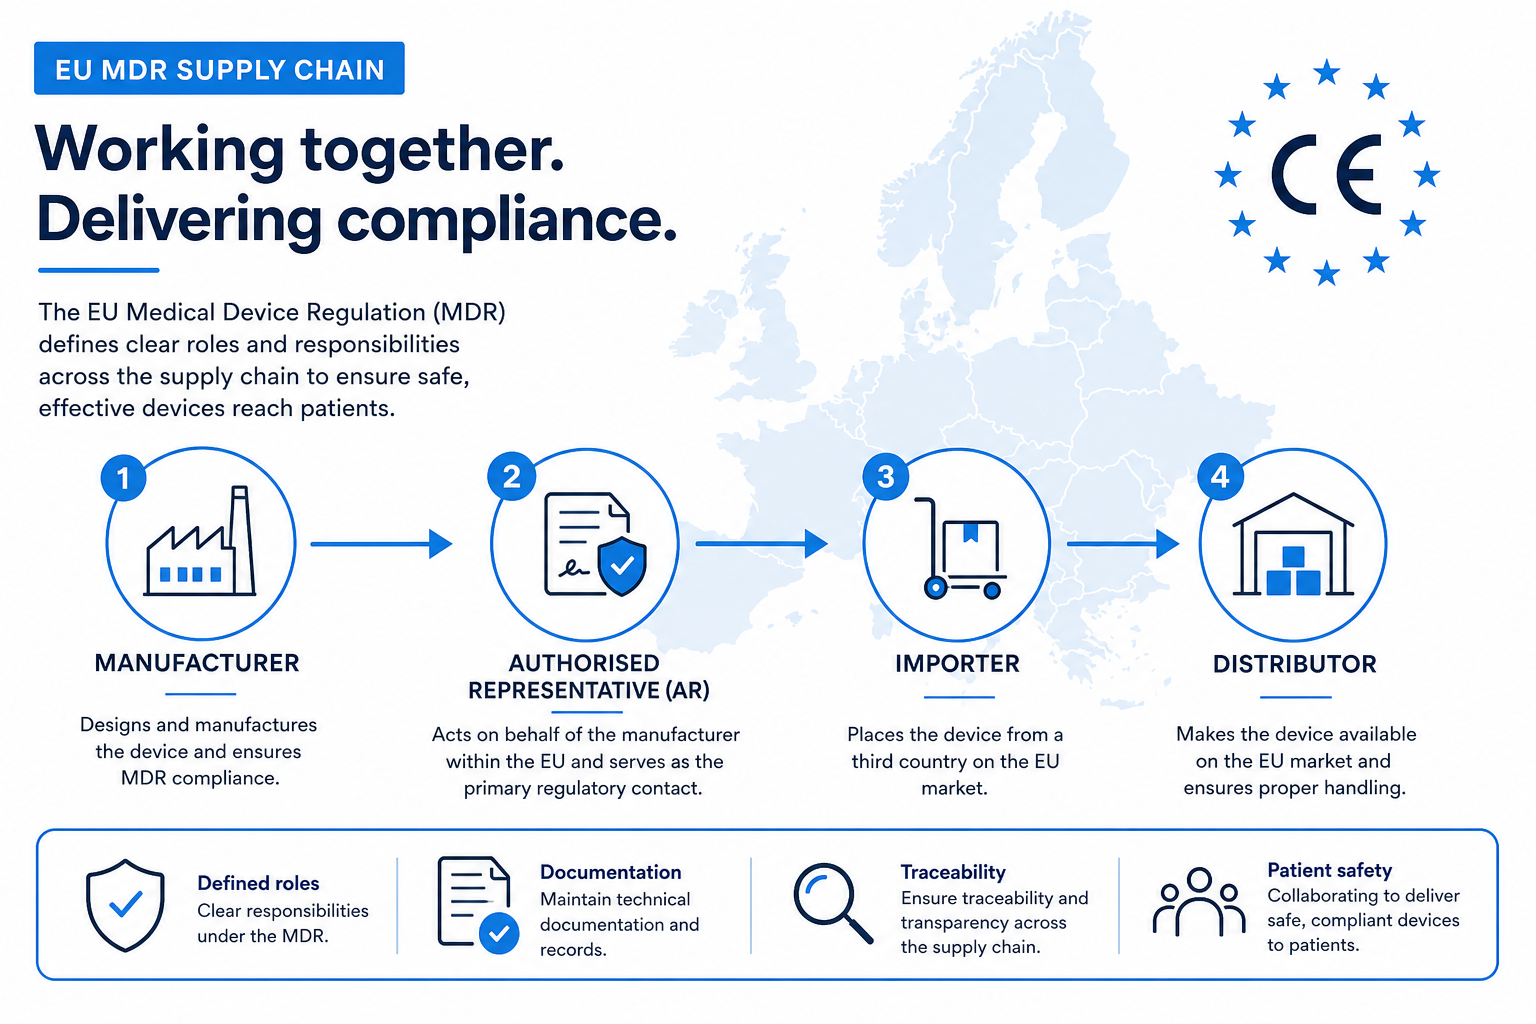
Short answer: The EU MDR (Regulation (EU) 2017/745) defines four "economic operators": the manufacturer (Article 10), the authorised representative / EC REP (Articles 11–12), the importer (Article 13) and the distributor (Article 14). Each has distinct legal obligations — responsibility does not sit with the manufacturer alone. Under Article 16, an importer or distributor that performs certain activities (placing a device on the market under its own name, changing the intended purpose, or some relabelling/repackaging) takes on all the obligations of a manufacturer.
Key takeaways
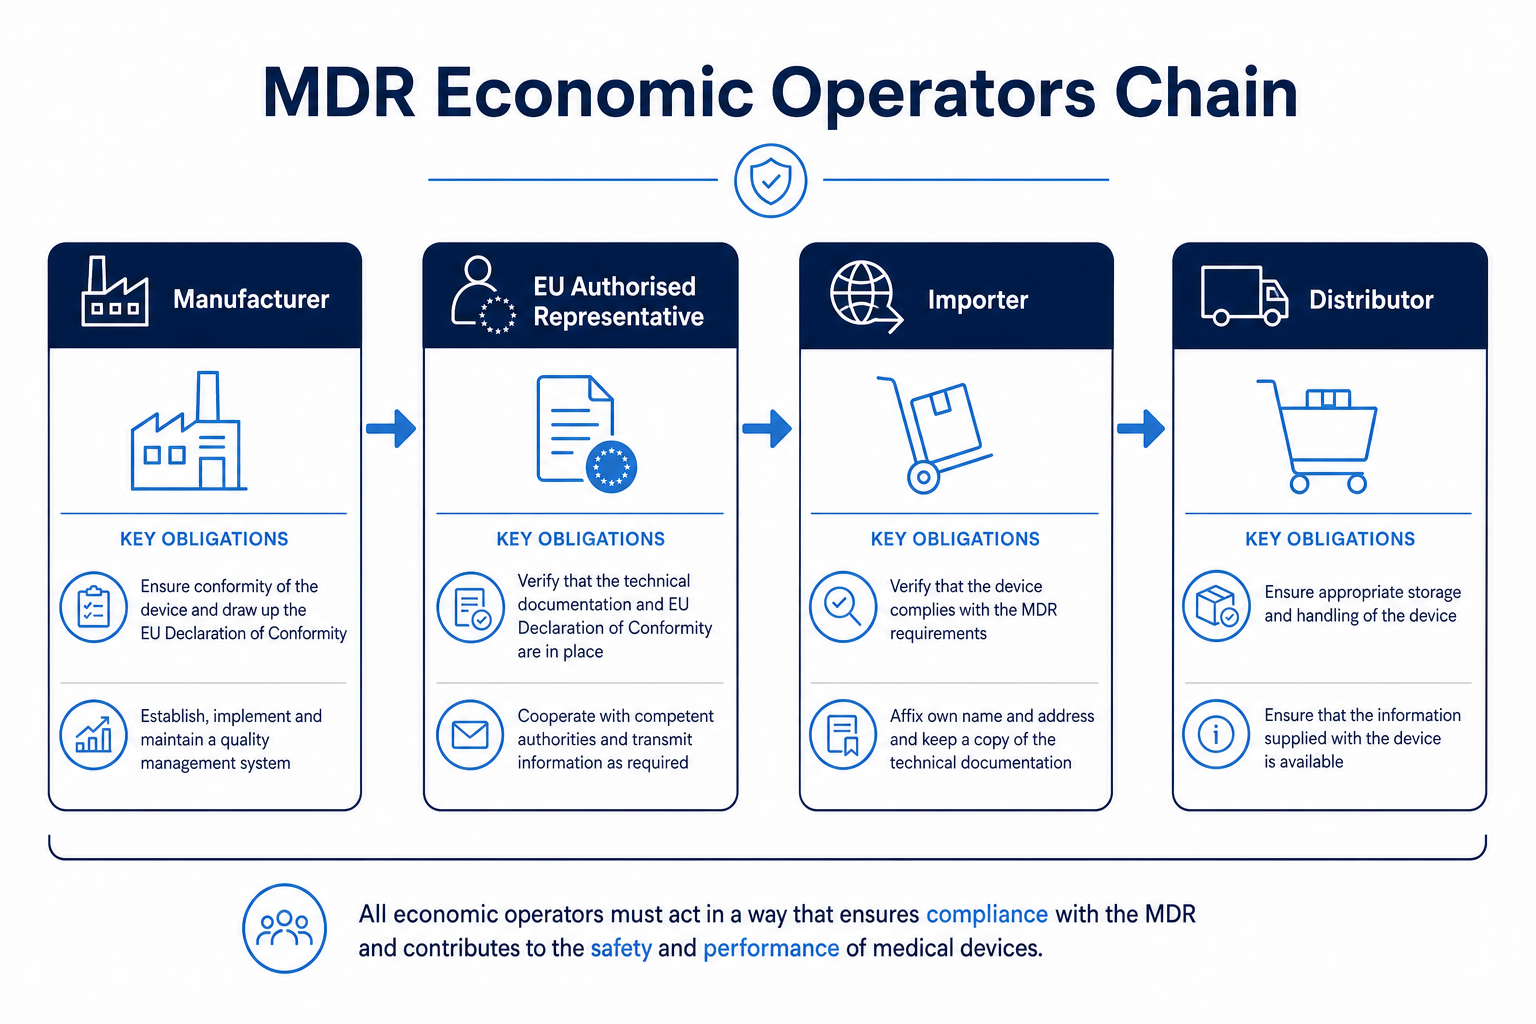
- Manufacturer: technical documentation, declaration of conformity, CE, PMS/PMCF, vigilance, PRRC, registration.
- Authorised representative: the EU-established legal contact appointed by a non-EU manufacturer under a written mandate; keeps the technical documentation available, liaises with authorities, has a PRRC, and can be jointly and severally liable in defined cases.
- Importer: places only compliant devices on the market; checks label/CE/registration/authorised representative, adds its own details, keeps records, forwards complaints and non-conformities.
- Distributor: before making a device available, checks the CE mark, EU declaration of conformity, label/IFU (including language), UDI and importer details; maintains transport/storage conditions; reports non-conformities.
- Article 16: draws the "who counts as a manufacturer" line — get it wrong and you inadvertently take on manufacturer obligations.
This article is part of the What is the EU MDR? pillar guide.
Manufacturer (Article 10)
The manufacturer is the natural or legal person that manufactures or fully refurbishes a device and markets it under its own name or trademark. Main obligations:
- Conformity with the applicable GSPR (Annex I) and a risk management system (ISO 14971).
- Preparing and maintaining the technical documentation (Annexes II–III).
- Conformity assessment, EU declaration of conformity, CE marking.
- A quality management system (ISO 13485-based).
- PMS/PMCF, PSUR, vigilance (serious incident and FSCA reporting), trend reporting.
- Assigning the UDI and registering in EUDAMED (ÜTS in Türkiye).
- Appointing a PRRC (Article 15).
- Adequate financial coverage for liability (defective devices).
- Providing documentation to authorities on request and carrying out corrective action.
Authorised representative — EC REP (Articles 11–12)
A manufacturer not established in the EU must appoint an EU-established authorised representative (by written mandate) to place a device on the EU market. The authorised representative:
- Keeps a copy of the technical documentation, DoC and certificates available in the EU.
- Responds to authority requests for information/samples and coordinates on corrective actions.
- Fulfils/verifies registration obligations (EUDAMED actor/device registration).
- Forwards vigilance reports; passes complaints to the manufacturer.
- Has its own PRRC.
- Can be jointly and severally liable with the manufacturer for defective devices where the manufacturer has breached its obligations.
The scope and termination conditions of the mandate must be clearly defined in the contract; a handover procedure applies when the representative changes.
Importer (Article 13)
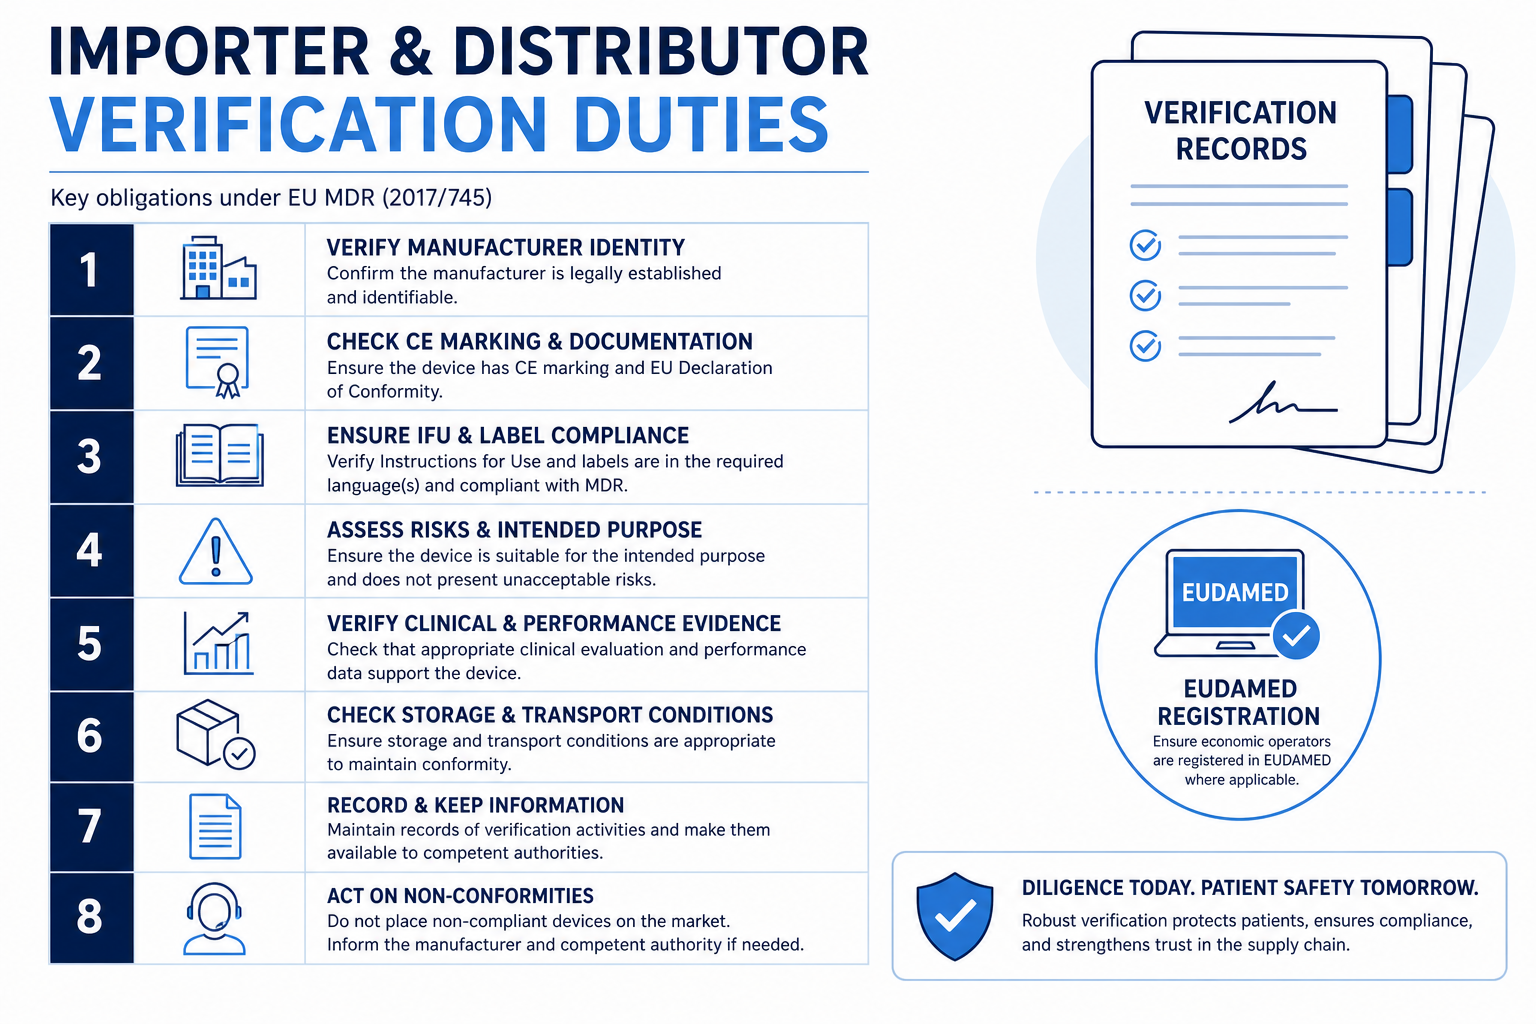
The EU-established party that first places a device from a third country on the EU market. Before placing it, the importer verifies:
- The device is CE marked and an EU declaration of conformity has been drawn up.
- An authorised representative has been appointed and its details are accessible.
- Labelling, IFU and — where required — language conditions are met.
- The UDI has been assigned; the device is registered in EUDAMED (where applicable).
In addition: indicates its own name and contact details on the label/packaging (without altering the device), maintains storage/transport conditions, keeps complaint and suspect-device records, does not place a non-compliant device on the market and notifies the authority/manufacturer where needed, and keeps records for at least 10 years (15 years for implantable devices).
Distributor (Article 14)
Any party in the supply chain, other than the importer, that makes a device available. Before making it available, the distributor checks (on an appropriate sampling basis):
- The CE mark and EU declaration of conformity are present.
- The label and IFU are provided; in the required language.
- The UDI is present where applicable.
- The importer has indicated its details on the label/packaging.
Also: maintains transport/storage conditions as specified by the manufacturer, does not supply a device it believes to be non-compliant and informs the manufacturer/authorised representative/importer and, where needed, the authority, participates in complaint/recall processes, and keeps records.
When does an operator become a "manufacturer"? (Article 16)
An importer, distributor or other person takes on the manufacturer's obligations where it:
- Makes a device available under its own name or trademark (private label / OEM).
- Changes the intended purpose of a device already on the market.
- Modifies a device in a way that can affect conformity.
However, the following activities alone do not make you a manufacturer (Article 16(2)), if the conditions are met: adding information without touching the device; changing the translation/information on the outer packaging; repackaging without opening the device itself (typically to adapt it to the market). An operator doing these must have a quality management system, notify the competent authority 28 days in advance, and obtain a notified body certificate for the relabelling/repackaging procedures.
Notes for Türkiye
- Türkiye-established operators register products and companies via the ÜTS (Product Tracking System); a separate EUDAMED obligation arises on the EU side.
- A Türkiye-based manufacturer exporting to the EU must appoint an EU-established authorised representative (Türkiye is not an EU member).
- For companies importing into Türkiye, the TİTCK's import authorisation/registration rules also apply.
Frequently asked questions
Can one company be both importer and distributor? Yes; the obligations for each role apply separately.
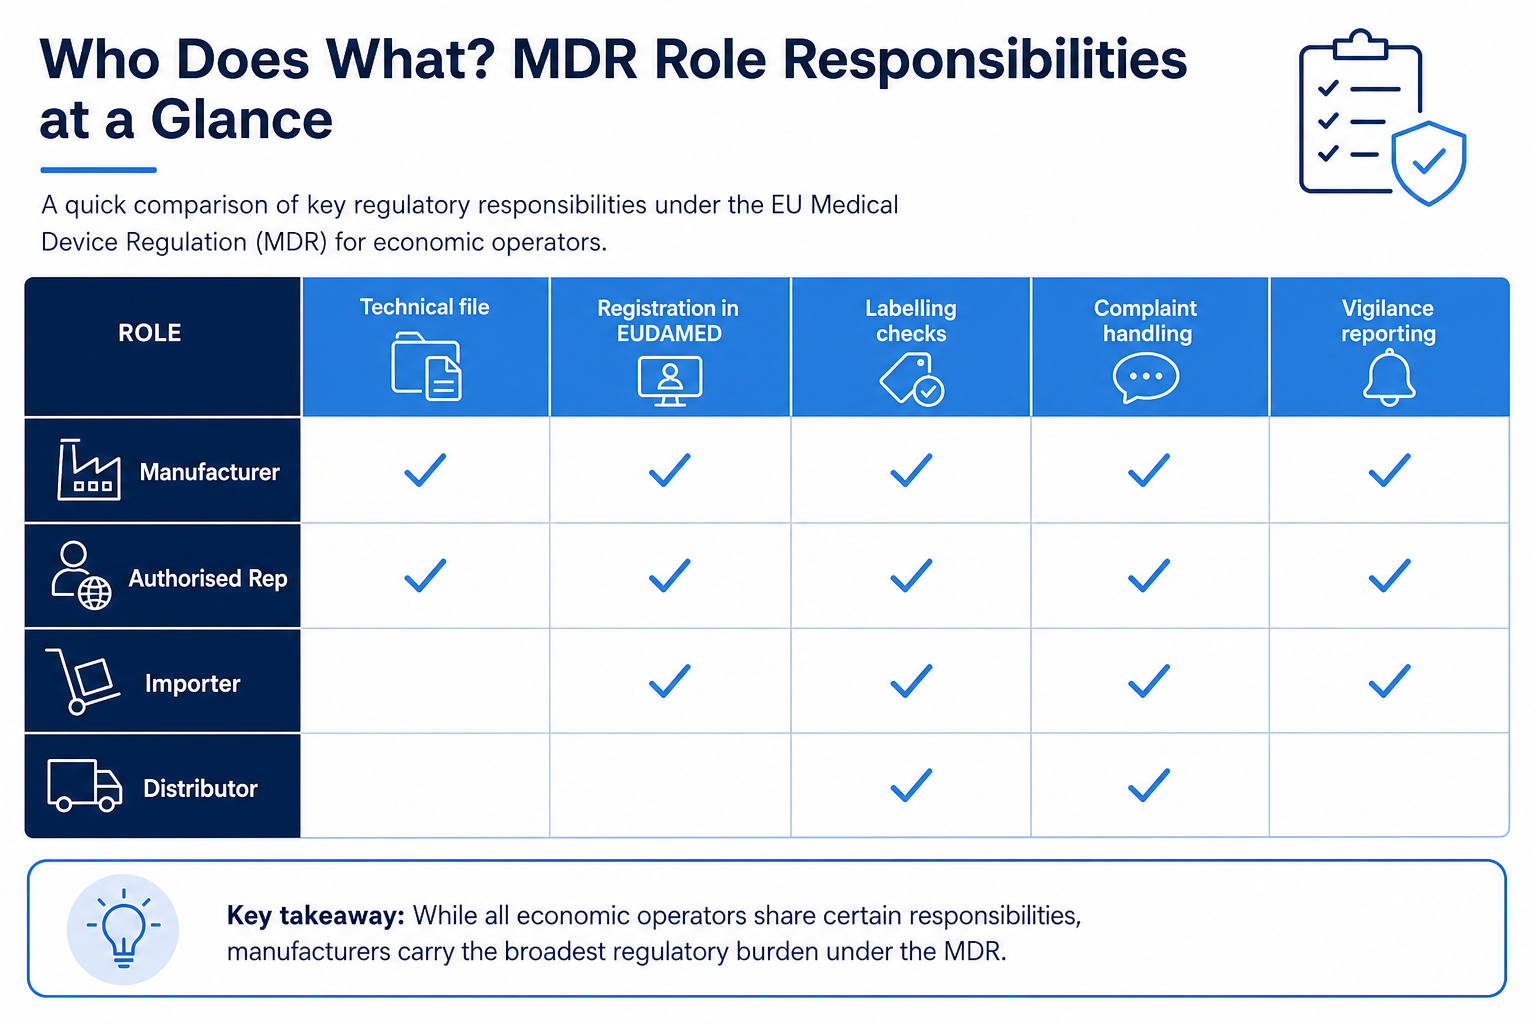
Does the authorised representative prepare the technical file? No; it does not prepare it but keeps it available and provides it to authorities. Preparation is the manufacturer's responsibility.
Does the distributor check every single device? No; checking on an appropriate sampling basis is sufficient, but the method must be documented.
What happens when I sell under my own private label? Under Article 16 you take on all the obligations of a manufacturer.
How long must I keep records? The general rule is 10 years; 15 years for implantable devices.
How MEGACERT helps
We support role determination, authorised representative contracts and handovers, importer/distributor verification procedures and Article 16 assessments; we also provide an EU Authorised Representative service. Let us clarify the roles in your supply chain in 30 minutes. → Contact. For the responsible person: Who is the PRRC? (cluster link)
Sources
- Regulation (EU) 2017/745, Articles 10–16 and Article 25 (traceability within the supply chain). EUR-Lex.
- MDCG 2021-27 — Questions and Answers on Articles 13 & 14 (importers and distributors).
- MDCG 2021-25 — legacy device obligations (including operator provisions).
- TİTCK. Product Tracking System (ÜTS) guides. https://www.titck.gov.tr
Last updated: 30 August 2026. General information only.